Biotechnology patent process and clinical trials
April 17, 2024
A conference abstract can create a filing deadline while a therapeutic candidate remains in preclinical development. Years later, new clinical findings can raise claim-support questions while an existing patent approaches a maintenance payment. For patent counsel and in-house IP teams, the challenge is coordinating these decisions without tying legal deadlines to clinical phases. A clear view of both processes helps you decide what to file, what evidence supports the claims, and which obligations need their own calendar.
Scope: U.S. utility patents and human therapeutic development, international filing, and European protection. Attorneys retain review and decision authority.
Two parallel processes with different decision-makers
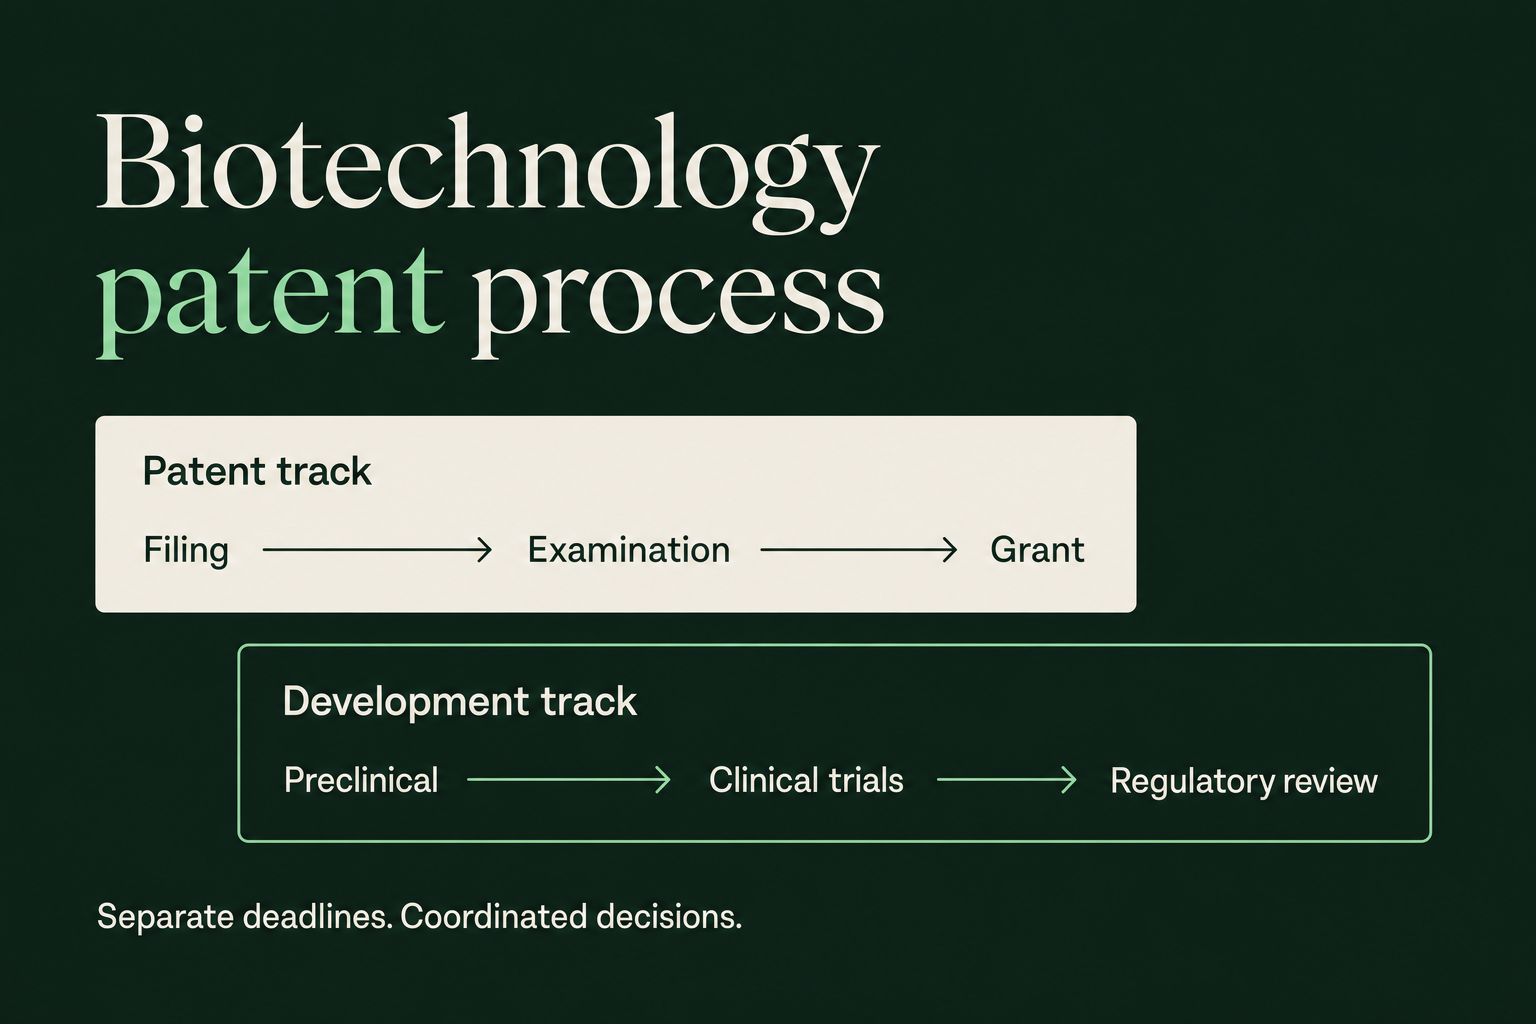
The biotechnology patent process concerns the invention and the claims that define its legal protection. The U.S. Patent and Trademark Office (USPTO) examines patentability. The Food and Drug Administration (FDA) oversees covered clinical investigations and reviews the appropriate marketing application. Each process has its own requirements and deadlines.
Keep separate records for:
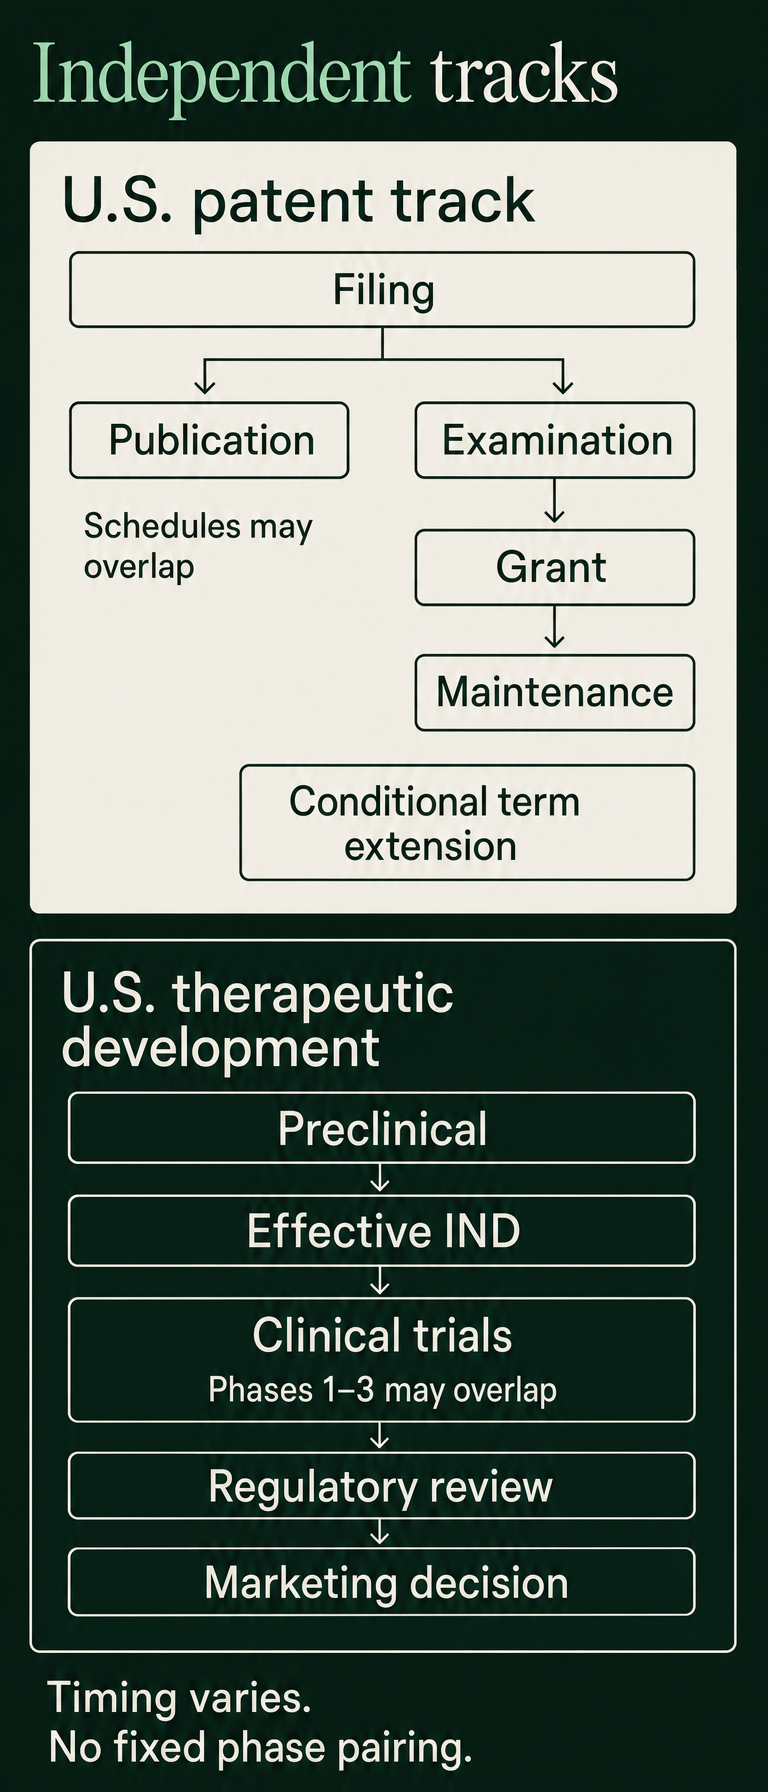
- The patent track: filing, application publication, examination, allowance, grant, maintenance, and any eligible term adjustment or extension.
- The development track: preclinical research, an effective Investigational New Drug (IND) application where required, clinical trials, and regulatory review.
Calendar overlap creates coordination work. It supplies no fixed pairing between a trial phase and a patent event. Patent grant depends on patent-office proceedings; clinical progression depends on the development evidence and regulatory requirements. USPTO process guidance and FDA investigation rules describe these separate processes.
Track 1: filing and prosecuting a U.S. biotechnology patent
1. Define the invention and its supporting disclosure
Start with the technical contribution: a composition, production process, formulation, or therapeutic method, for example. Gather the inventors' contributions, experimental methods, results, alternatives, and planned public disclosures. Counsel assesses patent eligibility, novelty, nonobviousness, utility, and disclosure sufficiency against the proposed claims.
A prior-art search reviews earlier patent documents and scientific literature against the proposed invention. Counsel uses relevant disclosures and dates to assess patentability and guide claim drafting.
A therapeutic patent filing can precede human trials. USPTO guidance recognizes that appropriately correlated laboratory or animal evidence can support therapeutic utility. Human clinical data and FDA approval are not prerequisites for that patent requirement. The filing decision still depends on what the application actually teaches. Therapeutic utility guidance separates the patent standard from FDA safety and efficacy review.
For biotechnology patent applications, claim breadth deserves particular attention. Enablement means teaching a skilled person to make and use the claimed invention without undue experimentation. Written description requires adequate disclosure of the claimed subject matter. These disclosure requirements make the filing record important: counsel should map the proposed scope to the experiments, examples, and technical teaching available at filing.
2. Choose the filing route and protect disclosure timing
A U.S. nonprovisional application enters the examination process. An optional provisional application can establish an earlier filing date for adequately disclosed subject matter. The provisional is unexamined and requires a further filing to pursue a patent. Normally, a corresponding nonprovisional claiming its benefit must be filed within 12 months. The earlier date applies only where the provisional supports the later claimed subject matter. Provisional filing requirements explain these conditions.
Give counsel the publication, presentation, collaboration, and commercial-disclosure calendar before releasing invention details. A pre-filing disclosure can jeopardize foreign patent rights even where a U.S. grace-period exception may apply. Clinical readiness and publication readiness therefore belong in separate filing discussions.
For international protection, the Patent Cooperation Treaty (PCT) provides an international application route. A PCT application claiming priority from an initial filing is generally filed within 12 months. National or regional phase entry is usually due around 30 months from priority, with office-specific deadlines and requirements. Grant decisions remain with the relevant national or regional offices. WIPO's PCT explanation describes this staged route.
3. Track application publication separately
U.S. utility applications generally publish 18 months from the earliest claimed filing or priority date, subject to exceptions. Publication makes the application available to the public. Publication and examination schedules can overlap. Grant follows a separate patentability decision and issuance process.
Record the publication number separately from any later patent number. Also distinguish application publication from the team's own journal paper or conference abstract. Those events can have different disclosure and prior-art consequences. The USPTO's application publication rules describe the general timing and nonpublication conditions.
4. Respond to examination using supported claims and evidence
During prosecution, the proceedings between the applicant and the patent office, an examiner may raise objections or reject claims. Counsel reviews the cited art and legal grounds, then chooses arguments, supported amendments, or further procedural steps. Responses have their own deadlines, regardless of whether the candidate is in Phase 1, Phase 2, or another development stage.
Later clinical findings require a specific distinction:
- Evidence about the disclosed invention: a later declaration may help demonstrate that the invention works. Enablement is assessed as of filing, and later evidence cannot supply an enabling disclosure that was missing then. USPTO enablement guidance addresses this boundary.
- A new claim limitation: counsel identifies support in the application as filed before proposing an amendment. New matter cannot simply be inserted into the existing disclosure. Written-description requirements govern the support analysis.
- A newly developed invention: a new formulation, manufacturing method, or other advance may warrant a separate filing. Counsel determines the filing route and the date each claimed invention can receive.
Treat a Phase 2 data package as an input to this analysis. Its clinical label gives it no automatic entitlement to broader claims or the original filing date.
5. Separate allowance from grant
A notice of allowance indicates that the application has reached the allowance stage. Patent issuance follows payment of the issue fee and completion of the remaining requirements. Record both events, then review the granted claims against the product and manufacturing roadmap. A granted patent gives its owner a right to exclude others from covered acts, as the USPTO grant explanation describes.
A patentability assessment asks whether claims can qualify for protection. Freedom to operate (FTO) addresses potential exposure to others' patent rights for a defined activity in particular jurisdictions. Keep the FTO review workflow separate from prosecution and regulatory review.
6. Maintain the granted patent within its term
For covered U.S. utility patents, maintenance fees are due at 3.5, 7.5, and 11.5 years after grant. A 6-month surcharge grace period follows each due date. Nonpayment can cause expiration. These payments maintain the patent within its available term; they add no fresh term. USPTO maintenance guidance sets out the payment windows.
The ordinary modern U.S. utility-patent term begins at grant and generally ends 20 years from the applicable nonprovisional or international filing date, including relevant earlier domestic-benefit filings. Provisional and foreign-priority dates generally do not start that calculation. Adjustments, extensions, disclaimers, and lapse can affect the actual expiration date. Counsel should calculate it from the patent's filing chain and record. Patent-term rules supply the baseline.
Conditional U.S. term mechanisms
Long clinical development makes remaining patent life a commercial concern. Separate the mechanisms that can affect it.
Patent term adjustment (PTA) under 35 U.S.C. §154(b) addresses specified USPTO delays. Statutory limits and applicant-delay reductions affect the result. Its basis is patent prosecution. PTA provisions define the qualifying delays.
Patent term extension (PTE) under 35 U.S.C. §156 concerns qualifying regulatory review of specified products. Eligible claims cover a qualifying product, its use, or manufacture. The patent must remain unexpired and satisfy the applicable first-marketing and prior-extension conditions. Approval creates a time-sensitive assessment: the ordinary application window is 60 days from permission for commercial marketing or use. PTE eligibility requirements and filing-window rules govern the request.
For modern patents, PTE is capped at 5 years. A 14-year post-approval limit can further reduce the extension, based on the patent term remaining after approval. The calculation includes qualifying post-grant review time, testing-period treatment, and diligence reductions, so the awarded period can be shorter. Extension calculation rules explain the limits. Counsel also maps the approved product and uses to the statutory extended-period rights.
For a patent approaching expiry while qualifying regulatory review remains pending, counsel should assess the separate interim-extension provisions early. Phase 3 entry itself triggers neither an extension nor a maintenance payment.
Track 2: preclinical studies, clinical trials, and FDA review
The development track gathers the evidence needed to study and potentially market a therapeutic product. Its stages serve different purposes:
- Preclinical research: laboratory and animal work assesses toxicity and informs whether human testing should proceed. FDA preclinical guidance describes this work.
- IND effectiveness: covered human investigations generally require an effective IND. It may become effective 30 days after FDA receipt if FDA has imposed no clinical hold, or earlier upon FDA notification. The sponsor must satisfy the applicable study requirements before human administration. FDA IND procedures describe this regulatory gate.
- Phase 1: initial human studies examine pharmacology, metabolism, dose-related side effects, and, where possible, early effectiveness. Participants may be patients or healthy volunteers.
- Phase 2: controlled studies evaluate effectiveness for the indication in patients with the condition and assess common short-term side effects and risks.
- Phase 3: expanded studies gather further effectiveness and safety information to evaluate benefit-risk and support labeling. The clinical-phase regulation permits phases to overlap.
- Marketing application and review: the sponsor submits the appropriate application for the product's legal classification. Covered biological products follow the biologics license application (BLA) framework. Drugs subject to the applicable drug-approval framework may use a new drug application (NDA).
Keep regulatory status, patent status, and FTO conclusions as separate fields in development reports. Each answers a different question about the candidate's path toward commercialization.
European patents and EU supplementary protection
European patent practice has its own calendar. A European Patent Office (EPO) application undergoes search, publication, substantive examination on request, and a grant or refusal decision. After grant, the team addresses national validation or requests unitary effect where available. EPO procedure guidance explains these steps.
Under the European Patent Convention (EPC), the baseline term is 20 years from filing. EPO application renewal fees start in the 3rd year. Post-grant renewal obligations follow the chosen protection route. EPC term provisions and application renewal provisions distinguish term from fee payments.
For qualifying medicinal products in the European Union (EU), a supplementary protection certificate (SPC) provides separate, product-limited protection taking effect when the basic patent's lawful term expires. Eligibility and duration depend on the patent and marketing authorization. The ordinary filing window is 6 months after authorization, or after patent grant if later. Duration is capped at 5 years, with a conditional 6-month pediatric extension. The EU medicinal SPC regulation supplies these rules. Handle SPC applications by jurisdiction, separately from U.S. PTE requests.
Fictional example: coordinating the tracks
Fictional example. All company, candidate, and event details are invented.
A biotech team develops candidate B-17 and a production method. It plans a conference abstract while preclinical work continues.
Before the abstract
Patent track: counsel evaluates the inventors' contributions and available experiments, narrows the proposed scope to the supported disclosure, and files a U.S. provisional before the planned release. The team records the 12-month follow-on deadline.
Development track: researchers continue preclinical studies and regulatory preparation. The filing decision and the decision to begin human testing have separate evidence requirements.
During clinical development
Patent track: after the follow-on filing, counsel receives an examiner's rejection. A new data package supports further evaluation of the disclosed method; it also reveals a different formulation. Counsel separates response evidence and supported amendments from the potential new formulation filing.
Development track: the clinical team evaluates Phase 2 findings and the next study decision. It sends the changed formulation, planned disclosures, and relevant results to IP counsel without treating clinical progress as patent allowance.
After grant and toward a marketing decision
Patent track: the docket tracks maintenance due dates from grant and expiration from the applicable filing chain. Counsel prepares an eligibility assessment for possible regulatory PTE before the marketing decision approaches.
Development track: the regulatory team maintains the submission and review record and promptly communicates any approval date. That handoff enables counsel to address an applicable extension deadline.
The shared work product is a coordination record: invention, product version, supporting evidence, jurisdiction, patent event, regulatory event, responsible owner, and next deadline. Patent and regulatory owners approve their respective entries.
Budget around legal deadlines and development decisions
For an in-house IP team, coordination has a practical business case:
- Cost: allocate follow-on filing, national-phase, prosecution, and maintenance spend to the jurisdictions and inventions that support the candidate and manufacturing roadmap.
- Speed: give counsel planned disclosure dates and usable experimental records early enough to prepare filings and responses before their deadlines.
- Roadmap alignment: reassess claim coverage when the formulation, indication, production method, or target market changes. Carry the decision and its rationale into the next development review.
Assign an owner to each legal deadline. A delayed study decision leaves those obligations on the calendar.
Where Patlytics fits
Use the relevant modules for the patent work generated by development decisions:
- Patent & NPL Search: search patent documents and non-patent literature (NPL), including PubMed, for candidate references.
- Invention Disclosures (IDF): turn inventor materials, slides, and spoken input into a reviewable invention disclosure form (IDF).
- Patent Drafting: turn reviewed invention disclosures and technical source materials into draft claims and application text.
- Chemical drafting: work with an embedded structure editor, chemical image-to-SMILES conversion, and ChemDraw input. SMILES (simplified molecular input line entry system) represents a molecular structure as text. Markush and R-group generation supports drafting chemical families through generic structures and defined variable substituents.
- Biological drafting: use sequence uploads and antibody complementarity-determining region (CDR) detection to prepare draft language.
- Claim Charting: chemical structures render within claim limitations and Invalidity and Infringement charts. Invalidity charts map candidate prior-art disclosures; Infringement charts map evidence about an investigated product.
- Office Actions: use the U.S. office action, pending claims, and specification for Rejection Mapping against affected claims and cited art. Use response strategy options, the auto invention summary, and propagated amendments to prepare a draft response.
Attorneys retain review and decision authority over all outputs.
See how Patlytics turns biotech disclosures and examination records into supported claims and response options for attorney review. Book a demo.
Biotechnology patent process and clinical trials
A conference abstract can create a filing deadline while a therapeutic candidate remains in preclinical development. Years later, new clinical findings can raise claim-support questions while an existing patent approaches a maintenance payment. For patent counsel and in-house IP teams, the challenge is coordinating these decisions without tying legal deadlines to clinical phases. A clear view of both processes helps you decide what to file, what evidence supports the claims, and which obligations need their own calendar.
Scope: U.S. utility patents and human therapeutic development, international filing, and European protection. Attorneys retain review and decision authority.
Two parallel processes with different decision-makers
The biotechnology patent process concerns the invention and the claims that define its legal protection. The U.S. Patent and Trademark Office (USPTO) examines patentability. The Food and Drug Administration (FDA) oversees covered clinical investigations and reviews the appropriate marketing application. Each process has its own requirements and deadlines.
Keep separate records for:
- The patent track: filing, application publication, examination, allowance, grant, maintenance, and any eligible term adjustment or extension.
- The development track: preclinical research, an effective Investigational New Drug (IND) application where required, clinical trials, and regulatory review.
Calendar overlap creates coordination work. It supplies no fixed pairing between a trial phase and a patent event. Patent grant depends on patent-office proceedings; clinical progression depends on the development evidence and regulatory requirements. USPTO process guidance and FDA investigation rules describe these separate processes.
Track 1: filing and prosecuting a U.S. biotechnology patent
1. Define the invention and its supporting disclosure
Start with the technical contribution: a composition, production process, formulation, or therapeutic method, for example. Gather the inventors' contributions, experimental methods, results, alternatives, and planned public disclosures. Counsel assesses patent eligibility, novelty, nonobviousness, utility, and disclosure sufficiency against the proposed claims.
A prior-art search reviews earlier patent documents and scientific literature against the proposed invention. Counsel uses relevant disclosures and dates to assess patentability and guide claim drafting.
A therapeutic patent filing can precede human trials. USPTO guidance recognizes that appropriately correlated laboratory or animal evidence can support therapeutic utility. Human clinical data and FDA approval are not prerequisites for that patent requirement. The filing decision still depends on what the application actually teaches. Therapeutic utility guidance separates the patent standard from FDA safety and efficacy review.
For biotechnology patent applications, claim breadth deserves particular attention. Enablement means teaching a skilled person to make and use the claimed invention without undue experimentation. Written description requires adequate disclosure of the claimed subject matter. These disclosure requirements make the filing record important: counsel should map the proposed scope to the experiments, examples, and technical teaching available at filing.
2. Choose the filing route and protect disclosure timing
A U.S. nonprovisional application enters the examination process. An optional provisional application can establish an earlier filing date for adequately disclosed subject matter. The provisional is unexamined and requires a further filing to pursue a patent. Normally, a corresponding nonprovisional claiming its benefit must be filed within 12 months. The earlier date applies only where the provisional supports the later claimed subject matter. Provisional filing requirements explain these conditions.
Give counsel the publication, presentation, collaboration, and commercial-disclosure calendar before releasing invention details. A pre-filing disclosure can jeopardize foreign patent rights even where a U.S. grace-period exception may apply. Clinical readiness and publication readiness therefore belong in separate filing discussions.
For international protection, the Patent Cooperation Treaty (PCT) provides an international application route. A PCT application claiming priority from an initial filing is generally filed within 12 months. National or regional phase entry is usually due around 30 months from priority, with office-specific deadlines and requirements. Grant decisions remain with the relevant national or regional offices. WIPO's PCT explanation describes this staged route.
3. Track application publication separately
U.S. utility applications generally publish 18 months from the earliest claimed filing or priority date, subject to exceptions. Publication makes the application available to the public. Publication and examination schedules can overlap. Grant follows a separate patentability decision and issuance process.
Record the publication number separately from any later patent number. Also distinguish application publication from the team's own journal paper or conference abstract. Those events can have different disclosure and prior-art consequences. The USPTO's application publication rules describe the general timing and nonpublication conditions.
4. Respond to examination using supported claims and evidence
During prosecution, the proceedings between the applicant and the patent office, an examiner may raise objections or reject claims. Counsel reviews the cited art and legal grounds, then chooses arguments, supported amendments, or further procedural steps. Responses have their own deadlines, regardless of whether the candidate is in Phase 1, Phase 2, or another development stage.
Later clinical findings require a specific distinction:
- Evidence about the disclosed invention: a later declaration may help demonstrate that the invention works. Enablement is assessed as of filing, and later evidence cannot supply an enabling disclosure that was missing then. USPTO enablement guidance addresses this boundary.
- A new claim limitation: counsel identifies support in the application as filed before proposing an amendment. New matter cannot simply be inserted into the existing disclosure. Written-description requirements govern the support analysis.
- A newly developed invention: a new formulation, manufacturing method, or other advance may warrant a separate filing. Counsel determines the filing route and the date each claimed invention can receive.
Treat a Phase 2 data package as an input to this analysis. Its clinical label gives it no automatic entitlement to broader claims or the original filing date.
5. Separate allowance from grant
A notice of allowance indicates that the application has reached the allowance stage. Patent issuance follows payment of the issue fee and completion of the remaining requirements. Record both events, then review the granted claims against the product and manufacturing roadmap. A granted patent gives its owner a right to exclude others from covered acts, as the USPTO grant explanation describes.
A patentability assessment asks whether claims can qualify for protection. Freedom to operate (FTO) addresses potential exposure to others' patent rights for a defined activity in particular jurisdictions. Keep the FTO review workflow separate from prosecution and regulatory review.
6. Maintain the granted patent within its term
For covered U.S. utility patents, maintenance fees are due at 3.5, 7.5, and 11.5 years after grant. A 6-month surcharge grace period follows each due date. Nonpayment can cause expiration. These payments maintain the patent within its available term; they add no fresh term. USPTO maintenance guidance sets out the payment windows.
The ordinary modern U.S. utility-patent term begins at grant and generally ends 20 years from the applicable nonprovisional or international filing date, including relevant earlier domestic-benefit filings. Provisional and foreign-priority dates generally do not start that calculation. Adjustments, extensions, disclaimers, and lapse can affect the actual expiration date. Counsel should calculate it from the patent's filing chain and record. Patent-term rules supply the baseline.
Conditional U.S. term mechanisms
Long clinical development makes remaining patent life a commercial concern. Separate the mechanisms that can affect it.
Patent term adjustment (PTA) under 35 U.S.C. §154(b) addresses specified USPTO delays. Statutory limits and applicant-delay reductions affect the result. Its basis is patent prosecution. PTA provisions define the qualifying delays.
Patent term extension (PTE) under 35 U.S.C. §156 concerns qualifying regulatory review of specified products. Eligible claims cover a qualifying product, its use, or manufacture. The patent must remain unexpired and satisfy the applicable first-marketing and prior-extension conditions. Approval creates a time-sensitive assessment: the ordinary application window is 60 days from permission for commercial marketing or use. PTE eligibility requirements and filing-window rules govern the request.
For modern patents, PTE is capped at 5 years. A 14-year post-approval limit can further reduce the extension, based on the patent term remaining after approval. The calculation includes qualifying post-grant review time, testing-period treatment, and diligence reductions, so the awarded period can be shorter. Extension calculation rules explain the limits. Counsel also maps the approved product and uses to the statutory extended-period rights.
For a patent approaching expiry while qualifying regulatory review remains pending, counsel should assess the separate interim-extension provisions early. Phase 3 entry itself triggers neither an extension nor a maintenance payment.
Track 2: preclinical studies, clinical trials, and FDA review
The development track gathers the evidence needed to study and potentially market a therapeutic product. Its stages serve different purposes:
- Preclinical research: laboratory and animal work assesses toxicity and informs whether human testing should proceed. FDA preclinical guidance describes this work.
- IND effectiveness: covered human investigations generally require an effective IND. It may become effective 30 days after FDA receipt if FDA has imposed no clinical hold, or earlier upon FDA notification. The sponsor must satisfy the applicable study requirements before human administration. FDA IND procedures describe this regulatory gate.
- Phase 1: initial human studies examine pharmacology, metabolism, dose-related side effects, and, where possible, early effectiveness. Participants may be patients or healthy volunteers.
- Phase 2: controlled studies evaluate effectiveness for the indication in patients with the condition and assess common short-term side effects and risks.
- Phase 3: expanded studies gather further effectiveness and safety information to evaluate benefit-risk and support labeling. The clinical-phase regulation permits phases to overlap.
- Marketing application and review: the sponsor submits the appropriate application for the product's legal classification. Covered biological products follow the biologics license application (BLA) framework. Drugs subject to the applicable drug-approval framework may use a new drug application (NDA).
Keep regulatory status, patent status, and FTO conclusions as separate fields in development reports. Each answers a different question about the candidate's path toward commercialization.
European patents and EU supplementary protection
European patent practice has its own calendar. A European Patent Office (EPO) application undergoes search, publication, substantive examination on request, and a grant or refusal decision. After grant, the team addresses national validation or requests unitary effect where available. EPO procedure guidance explains these steps.
Under the European Patent Convention (EPC), the baseline term is 20 years from filing. EPO application renewal fees start in the 3rd year. Post-grant renewal obligations follow the chosen protection route. EPC term provisions and application renewal provisions distinguish term from fee payments.
For qualifying medicinal products in the European Union (EU), a supplementary protection certificate (SPC) provides separate, product-limited protection taking effect when the basic patent's lawful term expires. Eligibility and duration depend on the patent and marketing authorization. The ordinary filing window is 6 months after authorization, or after patent grant if later. Duration is capped at 5 years, with a conditional 6-month pediatric extension. The EU medicinal SPC regulation supplies these rules. Handle SPC applications by jurisdiction, separately from U.S. PTE requests.
Fictional example: coordinating the tracks
Fictional example. All company, candidate, and event details are invented.
A biotech team develops candidate B-17 and a production method. It plans a conference abstract while preclinical work continues.
Before the abstract
Patent track: counsel evaluates the inventors' contributions and available experiments, narrows the proposed scope to the supported disclosure, and files a U.S. provisional before the planned release. The team records the 12-month follow-on deadline.
Development track: researchers continue preclinical studies and regulatory preparation. The filing decision and the decision to begin human testing have separate evidence requirements.
During clinical development
Patent track: after the follow-on filing, counsel receives an examiner's rejection. A new data package supports further evaluation of the disclosed method; it also reveals a different formulation. Counsel separates response evidence and supported amendments from the potential new formulation filing.
Development track: the clinical team evaluates Phase 2 findings and the next study decision. It sends the changed formulation, planned disclosures, and relevant results to IP counsel without treating clinical progress as patent allowance.
After grant and toward a marketing decision
Patent track: the docket tracks maintenance due dates from grant and expiration from the applicable filing chain. Counsel prepares an eligibility assessment for possible regulatory PTE before the marketing decision approaches.
Development track: the regulatory team maintains the submission and review record and promptly communicates any approval date. That handoff enables counsel to address an applicable extension deadline.
The shared work product is a coordination record: invention, product version, supporting evidence, jurisdiction, patent event, regulatory event, responsible owner, and next deadline. Patent and regulatory owners approve their respective entries.
Budget around legal deadlines and development decisions
For an in-house IP team, coordination has a practical business case:
- Cost: allocate follow-on filing, national-phase, prosecution, and maintenance spend to the jurisdictions and inventions that support the candidate and manufacturing roadmap.
- Speed: give counsel planned disclosure dates and usable experimental records early enough to prepare filings and responses before their deadlines.
- Roadmap alignment: reassess claim coverage when the formulation, indication, production method, or target market changes. Carry the decision and its rationale into the next development review.
Assign an owner to each legal deadline. A delayed study decision leaves those obligations on the calendar.
Where Patlytics fits
Use the relevant modules for the patent work generated by development decisions:
- Patent & NPL Search: search patent documents and non-patent literature (NPL), including PubMed, for candidate references.
- Invention Disclosures (IDF): turn inventor materials, slides, and spoken input into a reviewable invention disclosure form (IDF).
- Patent Drafting: turn reviewed invention disclosures and technical source materials into draft claims and application text.
- Chemical drafting: work with an embedded structure editor, chemical image-to-SMILES conversion, and ChemDraw input. SMILES (simplified molecular input line entry system) represents a molecular structure as text. Markush and R-group generation supports drafting chemical families through generic structures and defined variable substituents.
- Biological drafting: use sequence uploads and antibody complementarity-determining region (CDR) detection to prepare draft language.
- Claim Charting: chemical structures render within claim limitations and Invalidity and Infringement charts. Invalidity charts map candidate prior-art disclosures; Infringement charts map evidence about an investigated product.
- Office Actions: use the U.S. office action, pending claims, and specification for Rejection Mapping against affected claims and cited art. Use response strategy options, the auto invention summary, and propagated amendments to prepare a draft response.
Attorneys retain review and decision authority over all outputs.
See how Patlytics turns biotech disclosures and examination records into supported claims and response options for attorney review. Book a demo.
The Premier AI-Powered Patent Platform
Reduce cycle times. Increase margins. Deliver winning IP outcomes.
Book a DemoThe Premier AI-Powered
Patent Platform
Reduce cycle times. Increase margins. Deliver winning IP outcomes.